
Characterization of the Active Site and Mechanism for 1,6-Hexanediol Production from Tetrahydrofuran-Dimethanol over Pt-Based Catalysts
Terminal diols such as 1,4-butanediol (1,4-BDO) and 1,6-hexanediol (1,6-HDO) are important building-block chemicals and find application in the synthesis of specialty chemicals and a variety of polymers, primarily in polyesters and polyurethanes, and polyamides such as nylon-6,6.1,2 Currently, 1,6-HDO is produced industrially from cylohexanone/cyclohexanol (KA oil) by oxidation with nitric acid to form adipic acid, followed by hydrogenation of the dimethoxy ester to yield 1,6-HDO. The attractiveness of the overall process is tempered by low conversions (4-8% for the oxidation of KA oil), difficult separations, the use of non-renewable fossil feedstocks, and emission of greenhouse gases, namely N2O.3 Various attempts to synthesize 1,6-HDO, as well as other alpha,omega-diols, from biomass resources have been reported. Using supported catalysts that contain precious metals with a reducible metal oxide such as Rh-ReOx,4-9 and Ir-ReOx,10,11 tetrahydropyran-2-methanol (THP2M), a biomass-derived feedstock,12 can be converted into 1,6-HDO with high selectivity. However, the high cost of the catalyst, in combination with a low productivity, limits the industrial applicability of those systems.13 More recently, it has been shown that 1,6-HDO can be produced from THP2M without the use of precious metals. However, 1,6-HDO selectivity only reached ~40%.14 Therefore, this route is not likely to be industrially applicable soon either.
In an effort to make the route to 1,6-HDO from biomass more industrially viable, our group has developed a complete reaction network producing 1,6-HDO from woody biomass.15 The early steps in this process convert biomass to tetrahydrofuran-dimethanol (THFDM) with high selectivity, and will likely be discussed elsewhere at this conference. In this study, we focus on the final steps to 1,6-HDO from THFDM, shown in Scheme 1.
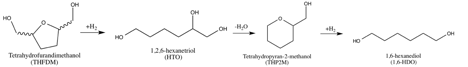
Scheme 1. Reaction sequence to 1,6-hexanediol (1,6-HDO) from tetrahydrofuran-dimethanol (THFDM)
We have developed a Pt-based catalyst that performs each step of the above reaction sequence with a total 1,6-HDO selectivity >90%. In comparison to the state-of-the-art, a previously described Rh-ReOx/C catalyst is not active for this reaction. Moreover, while Rh-ReOx/C is active for 1,6-HDO production from THP2M with selectivity >90%, the reaction rate is an order of magnitude lower than the total production rate of 1,6-HDO from THFDM over our Pt-based catalysts. Our catalyst, therefore, produces 1,6-HDO from a further upstream product, and at a much higher rate than the state-of-the-art Rh-ReOx/C system.
In situ characterization of this catalyst has revealed important information with regards to the active site and mechanism for 1,6-HDO production over our Pt-based catalysts. Our primary findings are that the oxidation state and acidity of the oxophilic additive (Mo, V, W, Re, for example) are crucial for this reaction. All catalysts are reduced with H2 prior to reaction, so the same conditions were used for characterization. We studied these catalysts with pyridine FTIR spectroscopy to obtain the Bronsted/Lewis acid ratio in each of our Pt-based catalysts, and compare that to reactivity data we had already compiled for the reaction of interest. As we observed that Bronsted acidity is correlated to the turnover frequency. Moreover, we found that this ratio increases in all catalysts during pre-reduction, a necessary step to achieve any reactivity. This data seems to indicate that the Bronsted acid site necessary for this reaction to take place, which are also confirmed by XPS and Raman characterizations. Furthermore, we are able to alter and characterize the distance between Pt nanoparticles and the oxophilic additive by changing the catalyst synthesis technique. Based on these studies we proposed a different mechanism to those which have been proposed previously, which will be discussed in more detail during the conference.
1 P. Werle, M. Morawietz, S. Lundmark, K. Sšrensen, E. Karivinen and J. Lehtonen, Alcohols, Polyhydric, in UllmannÕs Encyclopedia of Industrial Chemistry, 2012.
2 W. Tianfu, M. S. Ide, M. R. Nolan, R. J. Davis and B. H. Shanks, Energy Environ. Focus, 2016, 5, 13Ð17.
3 S. Van de Vyver and Y. Roman-Leshkov, Catal. Sci. Technol., 2013, 3, 1465Ð1479.
4 M. Chia, Y. J. Pag‡n-Torres, D. Hibbitts, Q. Tan, H. N. Pham, A. K. Datye, M. Neurock, R. J. Davis and J. A. Dumesic, J. Am. Chem. Soc., 2011, 133, 12675Ð12689.
5 M. Chia, B. J. OÕNeill, R. Alamillo, P. J. Dietrich, F. H. Ribeiro, J. T. Miller and J. A. Dumesic, J. Catal., 2013, 308, 226Ð236.
6 P. U. Karanjkar, S. P. Burt, X. Chen, K. J. Barnett, M. R. Ball, M. D. Kumbhalkar, J. B. Miller, I. Hermans, J. A. Dumesic and G. W. Huber, Catal. Sci. Technol., 2016, 6, 7841Ð7851.
7 Y. Kojima, S. Kotani, M. Sano, T. Suzuki and T. Miyake, J. Jpn. Pet. Inst., 2013, 56, 133Ð141.
8 K. Chen, S. Koso, T. Kubota, Y. Nakagawa and K. Tomishige, ChemCatChem, 2010, 2, 547Ð555.
9 Y. Nakagawa, M. Tamura and K. Tomishige, ACS Catal., 2013, 3, 2655Ð2668.
10 S. Koso, I. Furikado, A. Shimao, T. Miyazawa, K. Kunimori and K. Tomishige, Chem. Commun., 2009, 15, 2035Ð2037.
11 Y. Nakagawa, Y. Shinmi, S. Koso and K. Tomishige, J. Catal., 2010, 272, 191Ð194.
12 B. Xiao, M. Zheng, X. Li, J. Pang, R. Sun, H. Wang, X. Pang, A. Wang, X. Wang and T. Zhang, Green Chem., 2016, 18, 2175Ð2184.
13 Z. J. Brentzel, K. J. Barnett, K. Huang, C. T. Maravelias, J. A. Dumesic, and G. W. Huber, ChemSusChem., 2017, 10, 1-6.
14 S. P. Burt, K. J. Barnett, D. J. McClelland, P. Wolf, J. A. Dumesic, G. W. Huber, and I. Hermans, Green Chem., 2017, 19, 1390-1398.
15 J. He, K. Huang, K. J. Barnett, S. Krishna, D. M. Alonso, Z. Brentzal, S. P. Burt, T. W. Walker, W. Banholzer, C. T. Maravelias, I. Hermans, J. A. Dumesic, and G. W. Huber, Faraday Discussions, 2017.
See more of this Group/Topical: Catalysis and Reaction Engineering Division