
Thiyl radical chemistry is of exceptional interest in biology and organic chemistry owing to the ubiquity of thiol containing moieties found in nature and the availability of a diverse range of alkyl and aryl thiols with similar reactivities. S-centered radicals undergo efficient, essentially barrierless compared to alkyl and alkoxy radicals, anti-Markovnikov addition into unsaturated X=C(R)R’bonds. Of particular interest is the thiyl radical reaction with alkenes, radical-mediated thiol-ene coupling (TEC), which is one of the so-called “click” reactions and has become a staple in polymerization and conjugation chemistry. TEC reactions may be performed under ambient conditions and without solvent, exhibiting rapid, often quantitative conversions, and an insensitivity to water and oxygen, and offers the ability for spatio-temporal control. Robustness of the thiol-ene reaction is amplified when used in concert with thiolate anion-mediated reactions with electron deficient alkenes (thiol-Michael coupling), isocyanates, thioesters, or epoxides to generate linear and network polymers with increasing compositional and topological complexity as these reactions may be performed orthogonally to thiol-ene reactive alkenes. In our present efforts, our group’s focus is on fabricating polymers and networks with unique properties through the synergistic incorporation of thiyl radical and thiolate chemistries, including: sequence-ordered, DNA-mimetic polymers, covalent adaptable networks, and low dispersity oligomers and block copolymers.
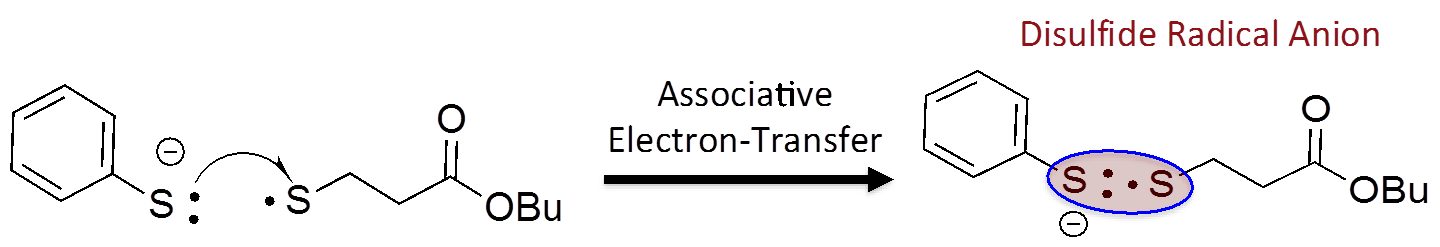
Despite being well regarded as insensitive to most chemical functionalities present in organic and biological systems, we observed significant retardation in TEC reaction and reduced conversions (<50%) when performed in the presence of amines, which were incorporated into the formulation either as the polymer side-group functionality (e.g. nucleobases), buffering system, or catalyst for thiolate reactions in dual-cure networks. Interested in the mechanism of this phenomenon, we investigated the impact of amines, and ammonium and thiolate ions on the reaction kinetics of TEC reactions using real-time Fourier transform infrared (rt-FTIR) spectroscopy in order to expand the guidelines for developing thiol-ene systems and to further understand the nature of S-centered radicals in the context of general synthetic chemistry. We determined that amines indirectly retard radical-mediated thiol-ene reactions through the generation of the thiolate anion in situ, which sequesters the catalytic reactivity of the thiyl radical through a radical—lone pair, two-sulfur three-electron (2S-3e) interaction, effectively partitioning the radical population into an intermediate disulfide radical anion (DRA) species. The degree of retardation increases proportionally with the ratio of thiyl to carbon radicals present under steady-state reaction conditions, the concentration of thiolate anion, and depends highly on the structure of the thiolate anion and the resulting disulfide radical anion.
Solution phase, density functional theory (DFT) calculations were employed to validate the mechanism by estimating, and comparing to experimental results, the retardation activities of various thiolate anions on the TEC kinetics. Computation revealed that all DRA structures possible in our systems are metastable with respect to the thiol-ene product and that the electron withdrawing character of the parent thiol substituent is the main factor in determining the DRA’s stability. Additionally, kinetic barriers to DRA formation were determined using parameters from Saveant theory on electron-transfer (ET) reactions, where the DRA formation was treated as a step-wise, associative ET reaction. Interestingly, the relatively high viscosity and low polarity/dielectric character of the reaction medium led to the solvent reorganization energy to be the more contributive parameter to the kinetic barrier in the formation than the theory previously assumed; therefore, thiolate anions with a more rigid structure and lower internal reorganization requirements displayed minimal barriers to DRA formation, which is in agreement with experimental data.
The phenomena of amine/thiolate induced retardation discussed in this paper has important implications for the design of thiol-ene systems when the reaction is performed under alkaline conditions in aqueous environments or in the presence of basic moieties in polymer/organic synthesis. Although computations and experiments showed that the DRAs are, in general, thermodynamically favorable to form, near quantitative conversions may be obtained by choice of the reacting thiol and/or alkene substrate, i.e. using a thiol with low acidity or an alkene, like norbornene or vinyl ether, which have rapid curing kinetics and are not rate-limited by the thiyl radical propagation step, like allyl ether and vinyl siloxane. Further, the importance of the solvent reorganization energy on the rate of ET highlights that conventional ET theories may not adequately describe ET reactions in solvent environments that polymer chemists frequently encounter (i.e. organic, weakly polar, and viscous).
See more of this Group/Topical: Materials Engineering and Sciences Division